Nanoparticle vaccines
Vaccination remains the most effective strategy for achieving durable protection against infectious diseases. In recent years, nanoparticle-based vaccines have emerged as a particularly promising approach, as they enable multivalent antigen presentation, improved lymphoid trafficking, and efficient activation of antigen-specific B cells. Nanoparticles can be constructed from a broad range of materials, including lipids, polymers, inorganic compounds, and self-assembling proteins. Their intrinsic physicochemical properties, combined with advances in genetic and biochemical engineering, allow for precise control over antigen density, orientation, and immune engagement.
The research focus of the Nanoparticle Vaccines group is to contribute to the rational design of next-generation vaccines by developing, characterizing, and preclinically evaluating B-cell-targeting nanoparticle platforms.
Self-assembling protein nanoparticles displaying native-like HIV-1 Env trimers
Self-assembling protein nanoparticles displaying native-like HIV-1 Env trimers have become a central strategy in HIV-1 vaccine design, as their repetitive antigen organization efficiently promotes B cell receptor crosslinking and activation of otherwise rare Env-specific B cell precursors. However, protein-based nanoparticle scaffolds can induce or recall scaffold-specific humoral immunity. Such responses may promote immunodominant activation of scaffold-specific memory B cells, accelerate particle clearance, and reduce antigen-focused B cell activation during homologous or heterologous boosting. This limitation is particularly relevant for biological scaffolds derived from licensed vaccines, such as human papillomavirus L1 virus-like particles (VLPs).
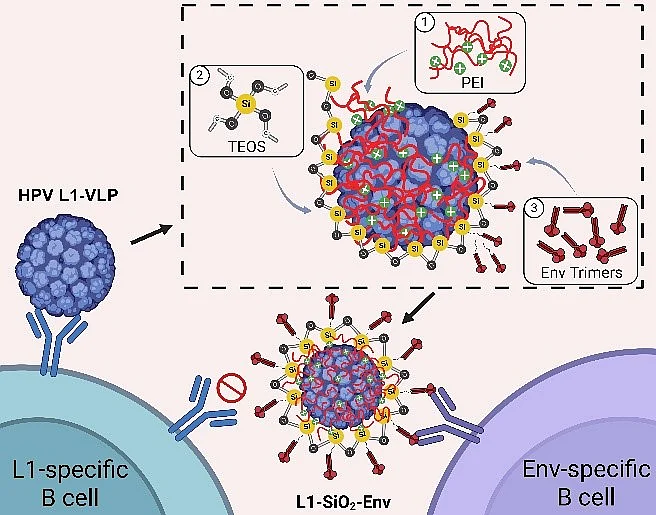
Currently, in cooperation with Prof. Matthias Epple, we designed a hybrid inorganic–protein nanoparticle platform in which self-assembling L1 VLPs are encapsulated within a thin silica shell to sterically shield L1-specific B cell epitopes. Silica shielding prevented recall of L1-specific memory B cells in HPV-preimmunized mice and abolished an anamnestic anti-L1 IgG response upon boosting. The silica shell further provided a chemically versatile surface for orthogonal antigen coupling. Using genetically encoded aldehyde tags in stabilized, native-like HIV-1 Env trimers previously designed in the lab, we were able to covalently attach these trimers via copper-free click chemistry to azide-functionalized silica surfaces.
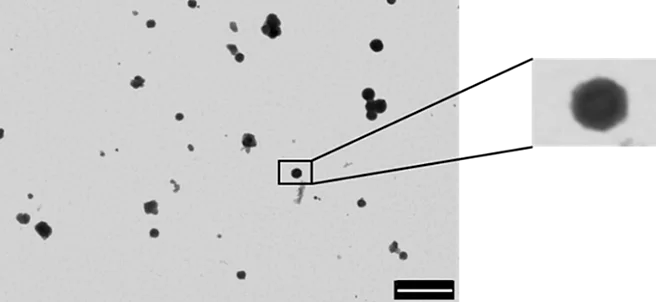
This strategy ensured high-density, repetitive presentation of conformationally intact Env trimers while maintaining the structural integrity of the nanoparticle. The resulting L1–SiO₂–Env nanoparticles induced robust, dose-dependent activation of Env-specific B cells in vitro. In vivo immunization experiments showed that L1–SiO₂–Env nanoparticles elicited strong Env-specific IgG responses while inducing only minimal L1-specific antibodies, confirming that scaffold shielding was maintained after antigen conjugation. Importantly, Env-specific antibody responses were enhanced rather than suppressed under conditions of pre-existing anti-HPV immunity, in contrast to non-shielded L1–Env nanoparticles, where recall of scaffold-specific immunity was associated with reduced Env-focused responses.
Together, these findings demonstrate that physical shielding of nanoparticle protein scaffolds enables control of scaffold-directed B cell recall and reshapes humoral immunodominance, thereby improving antigen-focused B cell responses in sequential vaccination settings. The silica-encapsulation strategy represents a modular approach to decouple scaffold immunogenicity from antigen presentation and can be extended to other protein-based nanoparticle platforms used in vaccine development.
Self-assembling lipid nanoparticles (LNPs)
Self-assembling lipid nanoparticles (LNPs) represent a clinically validated platform for nucleic acid delivery and have recently emerged as versatile tools for vaccine development. LNPs enable rational design of antigen display and intracellular delivery of genetic cargo. Induction of durable, high-affinity antibody responses critically depends on coordinated T follicular helper (TfH) cell differentiation and sustained germinal center (GC) reactions. However, currently available vaccine formulations do not allow selective modulation of antigen-activated B cells at the site of cognate T–B cell interactions. This limitation is particularly relevant for HIV-1 vaccine development, where germline-targeting strategies successfully prime rare precursor B cells but fail to sustain the prolonged GC activity required for broadly neutralizing antibody development.
Our central concept is the co-delivery of conformationally stabilized HIV-1 Env trimers on the LNP surface together with mRNA encoding genetic adjuvants. Antigen display is intended to drive BCR-mediated uptake by cognate B cells, while intracellular expression of immunomodulators provides spatially confined modulation of early TfH–B cell interactions within lymphoid tissues.
Our previous work with Env-conjugated liposomes, demonstrating BCR-specific targeting and activation of Env-specific B cells in vitro and in vivo, provides the technical foundation for generating Env-displaying LNPs capable of BCR-mediated uptake and intracellular delivery of immunomodulatory mRNA. The ongoing work includes optimization of mRNA composition for efficient expression in primary murine B cells, adjustment of lipid composition to minimize off-target uptake, evaluation of mRNA distribution in proliferating daughter B cells, and in vivo analysis of GC B cell and TfH responses following immunization with Env-conjugated, B cell–targeted LNPs.
In parallel, we performed initial screening of potential immunomodulatory candidates. Preliminary data demonstrate that genetic delivery of soluble checkpoint inhibitor constructs modulate vaccine-induced immune responses and enhances TfH frequencies in vivo. In addition, subtype-specific analyses of murine IFNα revealed marked heterogeneity in its capacity to promote B cell activation, GC-phenotype acquisition, and survival without impairing proliferation. These findings provide a mechanistic rationale for targeted, localized delivery of selected immunomodulators directly into antigen-activated B cells rather than systemic administration.
Together, this work establishes a precision-oriented vaccination strategy that integrates selected immunomodulators with antigen-specific B cell targeting. By combining multivalent antigen display with mRNA technology, the project provides a preclinical framework for selectively enhancing germinal center reactions and improving the quality of antibody responses against structurally complex viral antigens.